 Skip to content


Pharmaceutical tube filling requires defined controls for fill accuracy, cleanliness, traceability, and container closure. Whether equipment meets the applicable requirements depends on the product, facility, quality system, and validation plan.
This guide explains common cGMP, contamination-control, cleaning, dosing, and qualification topics. Regulatory expectations vary by jurisdiction and product; use the article as an equipment-selection checklist, not as regulatory approval.
The Foundations of Trust: Adhering to Global cGMP and FDA Standards
Current Good Manufacturing Practice (cGMP) regulations establish the baseline requirements for pharmaceutical manufacturing equipment. These standards exist to protect patient safety by preventing contamination, mix-ups, and errors that could compromise drug efficacy or introduce harmful substances into medications.
Non-Negotiable Machine Requirements for Cleanroom Environments
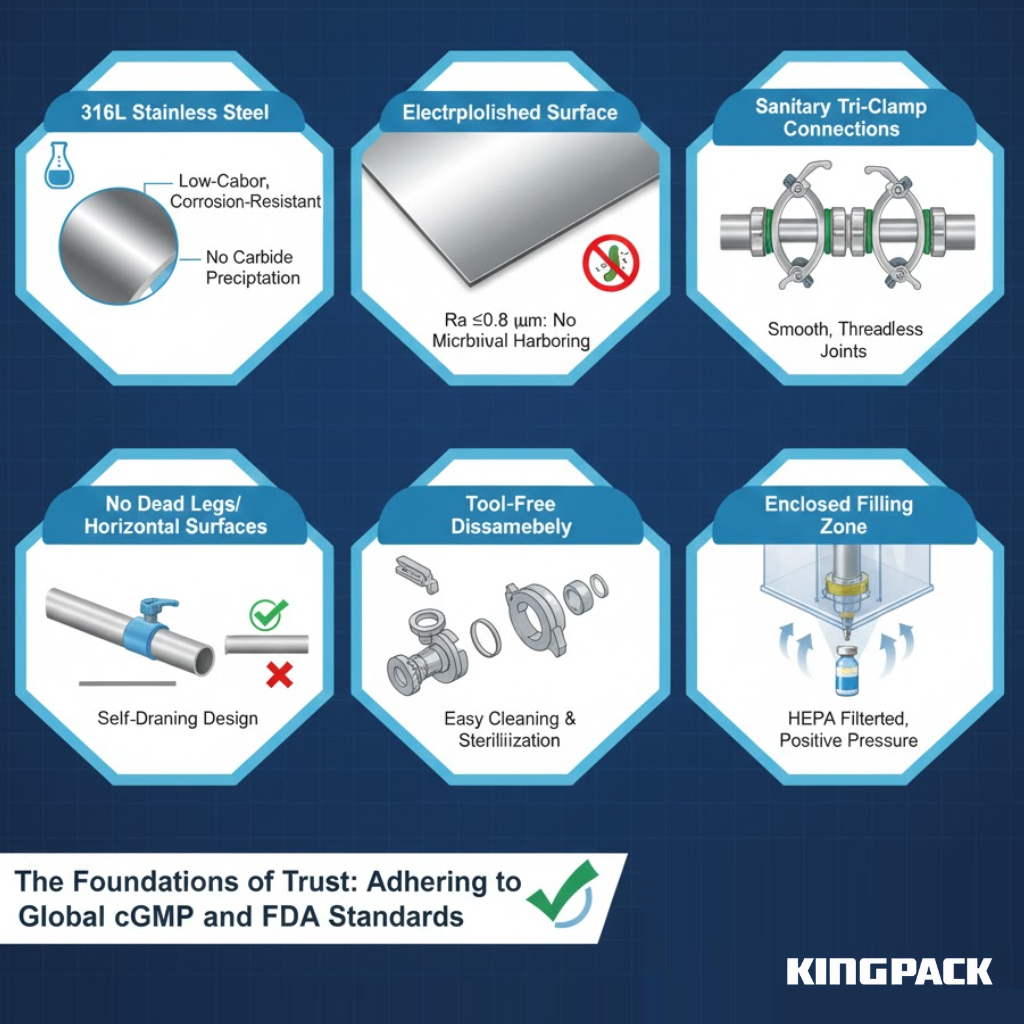
Pharmaceutical tube-filling equipment should support cleaning, contamination control, documentation, and the customer’s validated process. The applicable requirements must be defined for the facility and product.
- 316L stainless steel is commonly specified for product-contact parts, but the grade, finish, weld quality, and chemical compatibility should be confirmed in the technical specification.
- Surface roughness and electropolishing requirements are project-specific. Confirm the specified Ra value, inspection method, and cleaning-validation evidence rather than assuming a universal value.
- Sanitary fittings such as tri-clamps may simplify cleaning where included. Confirm gasket material, dead-leg design, drainability, and the supplied piping configuration.
- The design should be assessed for drainability, residue retention, and microbial-control risk. Do not assume every surface is dead-leg-free without drawings and inspection records.
- Tool-less removal may be available for selected nozzles, crimping tools, and closure parts. Confirm the actual disassembly procedure, cleaning method, and reassembly checks.
- Enclosures, airflow, and pressure control depend on the selected line and facility. Confirm the contamination-control strategy and qualification evidence for any HEPA or cleanroom features.
- Data logging, alarms, and recipe functions vary by control system. Confirm which parameters, records, users, and audit-trail features are included and how they will be validated.
FDA, EU GMP Annex 1, and other requirements are not interchangeable. Define the applicable jurisdiction, product, facility, and documentation responsibilities for the project.
Recommended Reading: What Is a Blister Pack? – Pharmaceutical Packaging Guide by King Pack – King Pack Machinery
Aseptic & Sterile Filling: Achieving Zero Contamination
Aseptic or sterile processing requires a validated contamination-control strategy. A single contaminated unit can be serious, but the risk and controls depend on the product, facility, container, and process.
Equipment may be configured to support aseptic or controlled filling, cleaning, and sterilization interfaces. The machine alone does not maintain sterility; the complete facility and quality system must be qualified.
Technology Deep Dive: CIP (Clean-in-Place) and SIP (Sterilization-in-Place) Automation
CIP or SIP automation may improve repeatability where the equipment is designed for it. Cycle parameters, coverage, chemistry, sterilization method, and documentation must be validated for the product.
Clean-in-Place (CIP) Systems
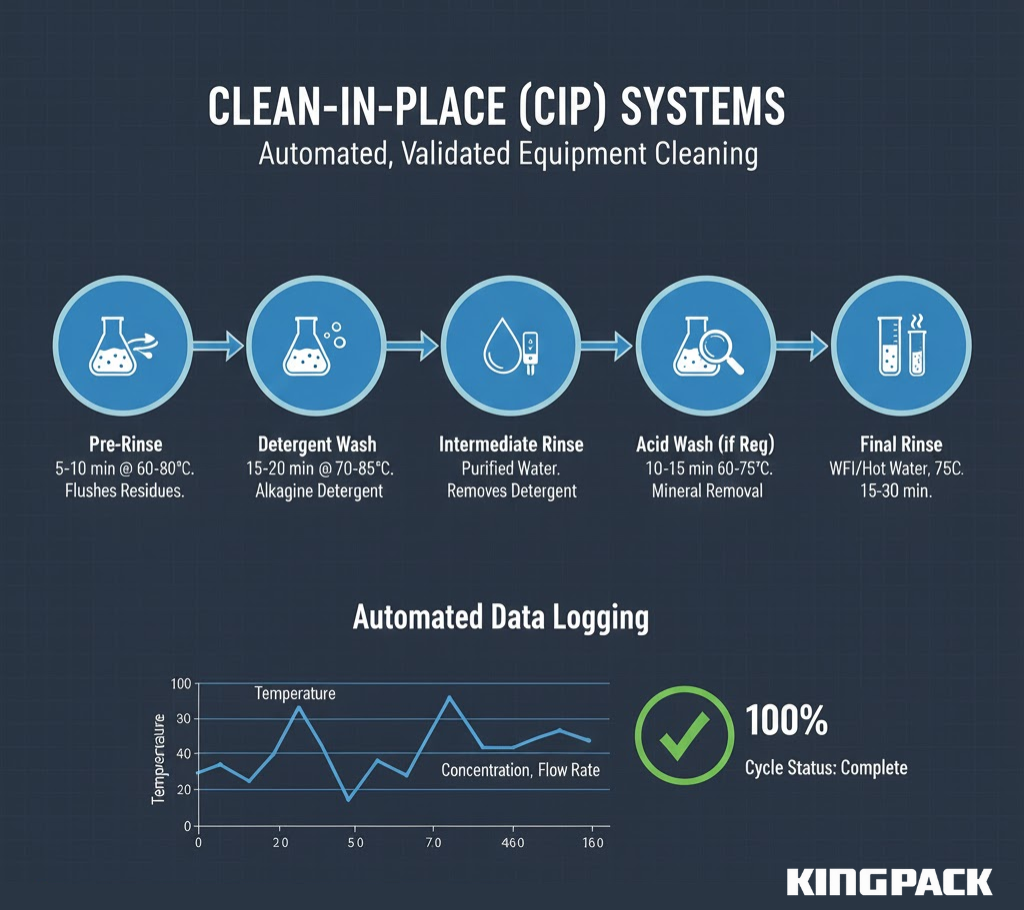
A CIP cycle is product- and equipment-specific. Typical stages, chemistry, temperature, time, flow, and acceptance criteria must be defined by the validated cleaning procedure.
- Pre-rinse parameters should be set by the validated cleaning procedure and product residue characteristics; do not apply a generic time or temperature.
- Detergent concentration, temperature, and contact time are formulation and equipment-specific and must be validated.
- Intermediate rinsing should continue until the validated residue or conductivity criteria are met.
- An acid wash may be used where justified by the residue and material compatibility; the chemistry and parameters require validation.
- Final-rinse water quality and acceptance criteria depend on the product, facility, and applicable pharmaceutical-water system.
- Sanitization parameters and method must be specified and validated for the equipment and process.
Automated monitoring can record cycle parameters and alarms. Acceptance criteria and deviation handling belong to the validated cleaning procedure.
Sterilization-in-Place (SIP) Systems
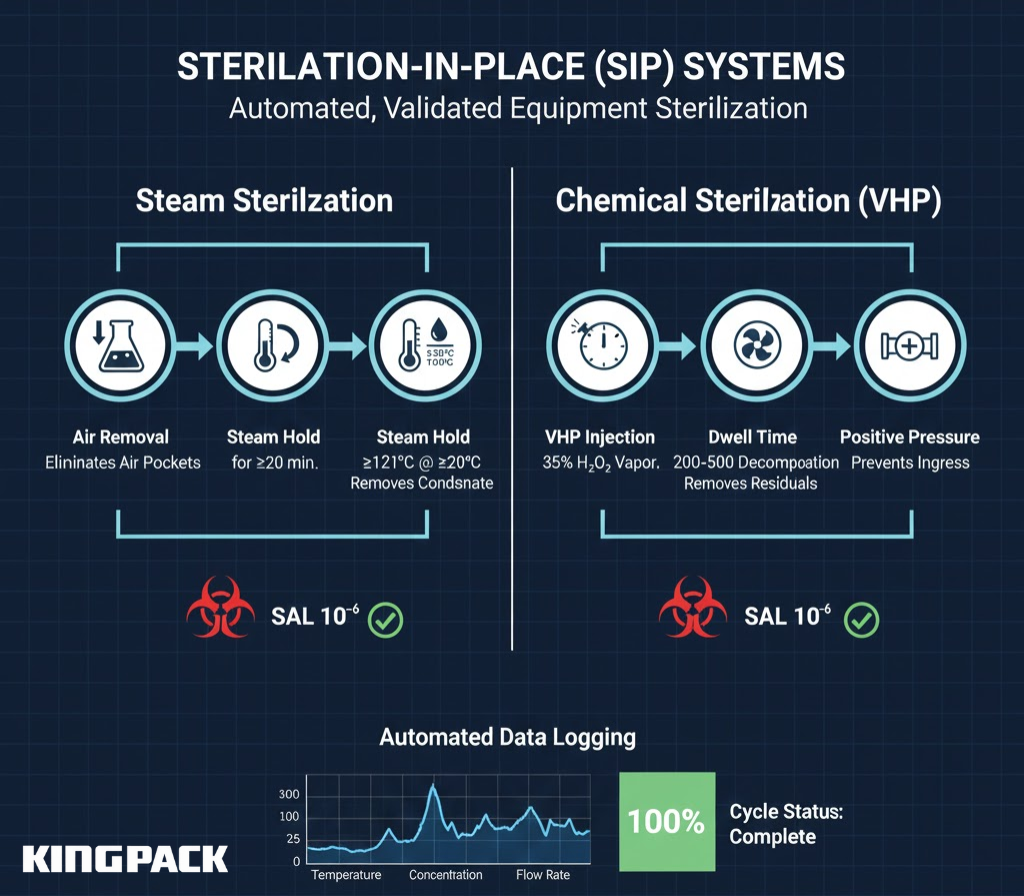
SIP performance and any sterility-assurance target must be established by the validated sterilization process. Do not treat a generic SAL value as a universal equipment claim.
Steam-sterilization temperature and hold time depend on the validated cycle, load, materials, and applicable standard.
- Remove all air pockets that would insulate surfaces from steam contact
- Maintain positive pressure to prevent contamination ingress during cooling
- Use condensate traps that remove water without allowing air entry
- Include temperature sensors at the coldest points to verify effective sterilization
VHP or other chemical sterilization may be suitable for compatible components, but concentration, exposure, aeration, and residual limits must be validated for the system.
Recommended Reading: World Top 10 Vial Filling Machine Manufacturers – King Pack Global Pharma Solutions – King Pack Machinery
Micro-Dosing Accuracy: Essential for High-Value APIs and Potent Ointments
Pharmaceutical dosing tolerance is product- and specification-specific. API cost and patient-safety considerations make calibration, in-process checks, and dose-uniformity testing important, but there is no universal accuracy percentage for every tube.
Pharmaceutical Dosing Standards
Acceptance limits should come from the product specification, pharmacopoeia, process capability, and applicable regulatory requirements.
| Product Type | Required Accuracy | Typical Technology | Validation Requirement |
| High-Potency Ointments | Example tolerance only; confirm the actual product specification and validated process capability. | Servo piston pumps | 100% weight verification |
| Standard Ointments | Example tolerance only; confirm the actual product specification and validated process capability. | Piston or gear pumps | Statistical process control |
| Gels and Creams | Example tolerance only; confirm the actual product specification and validated process capability. | Volumetric fillers | Periodic weight checks |
For high-potency or micro-dose products, small fill deviations can matter. The acceptable window must be calculated from the formulation, target dose, container, and validated measurement system.
Technologies for Ultra-Precise Dosing
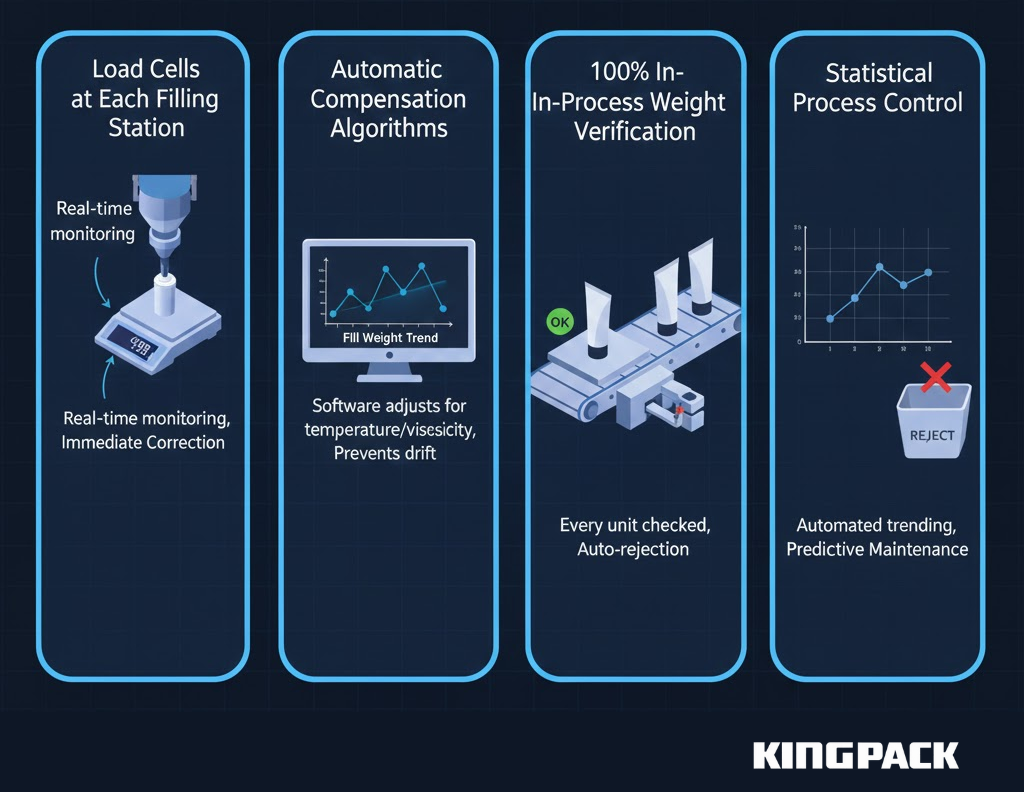
Positive-displacement, servo, peristaltic, or other dosing technologies may be selected according to the product and required control strategy.
- In-line weight monitoring may be available in selected configurations. Confirm sampling, calibration, response, and rejection logic.
- Automatic compensation may be available in some control systems, but its inputs, limits, and validation must be confirmed.
- 100% checkweighing and rejection are optional process features, not universal requirements. Confirm the actual inspection and reject system.
- Statistical process control can help detect drift when sufficient data and quality procedures are available.
Actual dosing capability must be demonstrated with representative product, tube, speed, and validated measurement methods; do not treat ±0.1% to ±0.2% as universal.
Recommended Reading: Types of Pharmaceutical Packaging – Comprehensive Guide by King Pack Packaging Solutions – King Pack Machinery
Validation Protocols: Your Blueprint for Machine Qualification
Validation demonstrates that the equipment and process can meet predefined requirements. The IQ/OQ/PQ approach, extent, and revalidation plan depend on the customer’s quality system and applicable regulations.
Structured Definition: The Roles of Installation Qualification (IQ) and Operational Qualification (OQ)
IQ, OQ, and PQ are commonly used qualification stages, but the exact protocol and responsibilities should be defined in the project validation plan.
Installation Qualification (IQ)
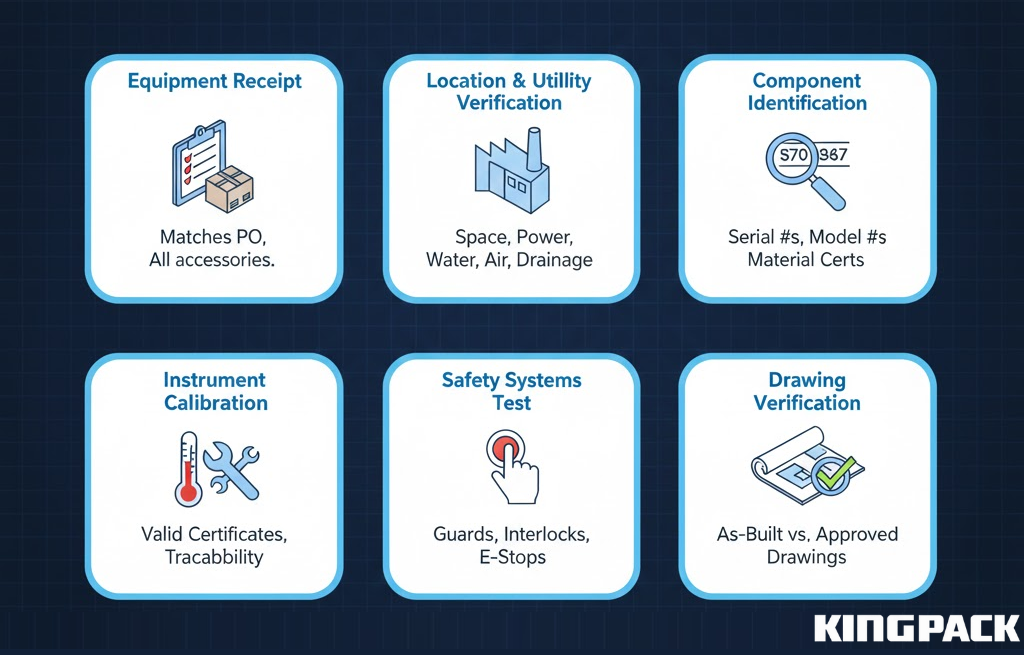
IQ verifies that equipment arrives complete and installs correctly according to specifications.
This phase documents:
- Equipment Receipt: Verification that delivered equipment matches purchase orders, including all specified options and accessories
- Location Verification: Confirmation that installation site provides adequate space, utilities, and environmental controls
- Utility Connections: Documentation of electrical connections, compressed air supply, water supplies, and drainage systems
- Component Identification: Recording serial numbers, model numbers, and material certificates for all equipment components
- Instrument Calibration: Verification that all measuring instruments (temperature sensors, pressure gauges, load cells) arrive with valid calibration certificates traceable to national standards
- Safety Systems: Testing of all guards, interlocks, and emergency stop functions
- Drawing Verification: Confirmation that as-built equipment matches approved engineering drawings
IQ documentation depth and test-point count vary by equipment, site, and quality system. Use the approved protocol and objective evidence.
Recommended Reading: Knowledge of Pharmaceutical Blister Packaging Machines – King Pack Packaging Solutions – King Pack Machinery
Operational Qualification (OQ)
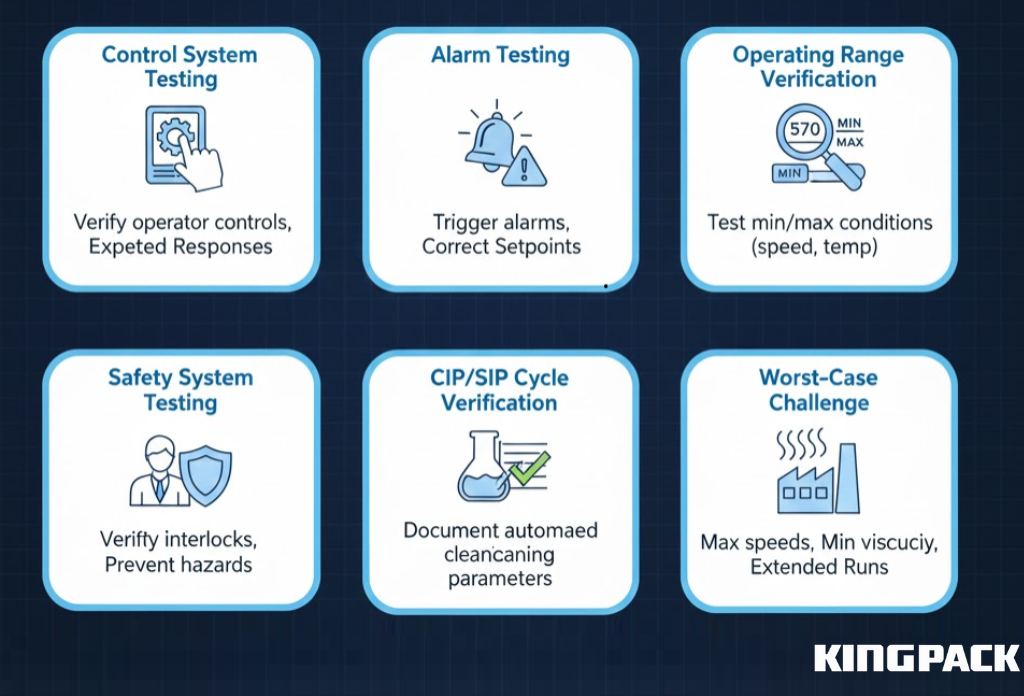
OQ tests the functions and operating ranges defined in the approved protocol, including controls, alarms, speeds, temperatures, and safety systems.
- Control System Testing: Confirms that PLC touch screens, control panels, and automated filling functions operate as designed, producing expected equipment responses.
- Alarm testing should cover the alarms and setpoints defined for the supplied equipment and risk assessment.
- Operating-range tests should use the approved minimum, nominal, and maximum conditions relevant to the product and line.
- Safety System Testing: Ensures tube elevators, tube holders, closure stations, and other safety interlocks prevent hazardous conditions during operation.
- CIP/SIP verification should demonstrate the approved cleaning or sterilization parameters for the actual equipment and product-contact path.
- Worst-case challenges should be justified by risk assessment and the approved qualification protocol.
OQ may use placebo or surrogate materials where appropriate, but the protocol determines the test material and acceptance criteria.
Performance Qualification (PQ): Documenting Consistent, Repeatable Production
PQ evaluates repeatable performance under defined production conditions using trained personnel, approved materials, and the customer’s quality procedures.
PQ Execution Requirements
The number of successful runs is protocol- and regulation-specific; three runs are common in some programs but should not be treated as a universal rule.
| PQ Requirement | Acceptance Criteria | Documentation |
| Fill Weight Uniformity | Fill-weight acceptance must follow the product specification and validated process capability. | Individual tube weights for entire run |
| Seal Integrity | Leak-test acceptance and sample size must be defined by the package and approved protocol. | Leak test results for minimum sample size |
| Batch Documentation | Complete, accurate records for all parameters | Batch production records with all signatures |
| Environmental Monitoring | All cleanroom parameters within specifications | Particle counts, microbial samples, temperature/humidity logs |
| Equipment Performance | OEE, downtime, and rejection targets should be set by the project and customer quality objectives, not a universal threshold. | Production logs showing speed, downtime, rejection rates |
Critical Process Parameters
PQ identifies and documents Critical Process Parameters (CPPs)—settings that significantly impact product quality. For tube filling, CPPs typically include:
- Fill volume setpoint and operating range
- Product temperature during filling
- Seal temperature and dwell time
- Line speed and indexing timing
- Nitrogen flush concentration (if used)
Changes to critical process parameters may require documented assessment and, where applicable, requalification or revalidation under the customer’s quality system.
Continued Process Verification (CPV)
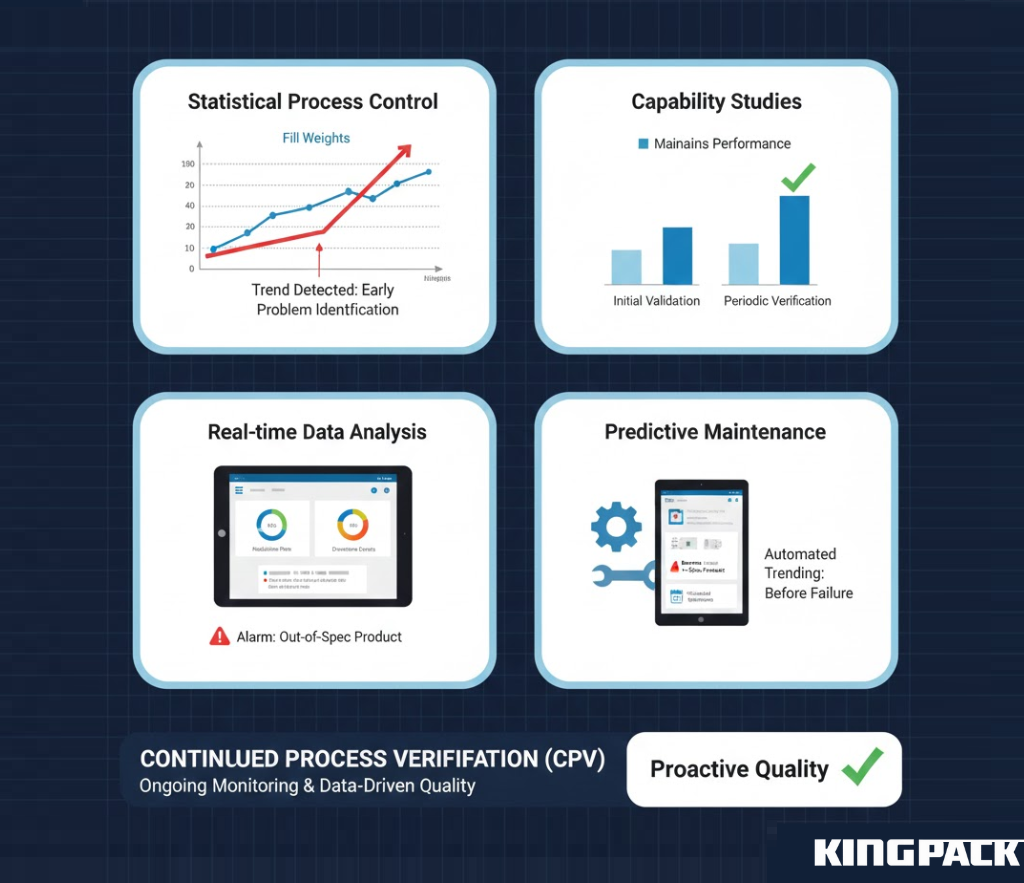
Continued process verification may use production data to monitor trends after qualification. The approach, metrics, and review frequency depend on the product and quality system.
Statistical process control charts track fill weights, rejection rates, downtime events, and other metrics. Periodic capability studies verify that equipment maintains validated performance levels. This data-driven approach provides earlier problem detection than periodic revalidation schedules.
Conclusion: Investing in Quality and Compliance for Your Pharma Future
Pharmaceutical tube-filling equipment should be specified against the applicable materials, contamination-control, documentation, and validation requirements. “Pharma-grade” is not a substitute for a project-specific assessment.
Qualification and validation provide evidence for the selected process, but CIP, SIP, dosing, and containment features must be evaluated within the complete facility and quality system.
Recommended Reading: Vial Filling Machines in the Pharmaceutical – King Pack Solutions for Precision & Sterility – King Pack Machinery
Request a Validated Consultation on Pharmaceutical Filling Equipment
King Pack can review the product, tube, fill range, hygiene, cleanroom, documentation, and validation requirements and then propose a project-specific configuration.
Frequently Asked Questions (Pharma FAQs)
What is the difference between Aseptic and Sterile filling in the pharmaceutical context?
Aseptic processing sterilizes or controls components separately before assembly in a controlled environment, while terminal sterilization sterilizes a sealed product after filling when the product and package tolerate it. The applicable approach depends on the product and validated process.
Why is Stainless Steel 316L mandatory for pharmaceutical contact parts?
316L is often selected for corrosion resistance and cleanability, but it is not universally mandatory. Material grade, finish, weld quality, and chemical compatibility should be specified for the actual process.
What are the three main stages of the machine validation process (IQ, OQ, PQ)?
IQ checks installation and documentation, OQ checks functions and operating ranges, and PQ checks repeatable performance under defined production conditions. The exact scope comes from the approved validation plan.
How do tube fillers prevent cross-contamination between different drug batches?
Cross-contamination controls may include cleaning validation, changeover procedures, product-contact design, line clearance, segregation, and batch records. The complete control strategy must be validated for the facility and products.
